
Mucus-targeting treatment for CF wins orphan drug status in Europe
Developer aiming to provide patients with 'improved therapeutic options'
Written by |
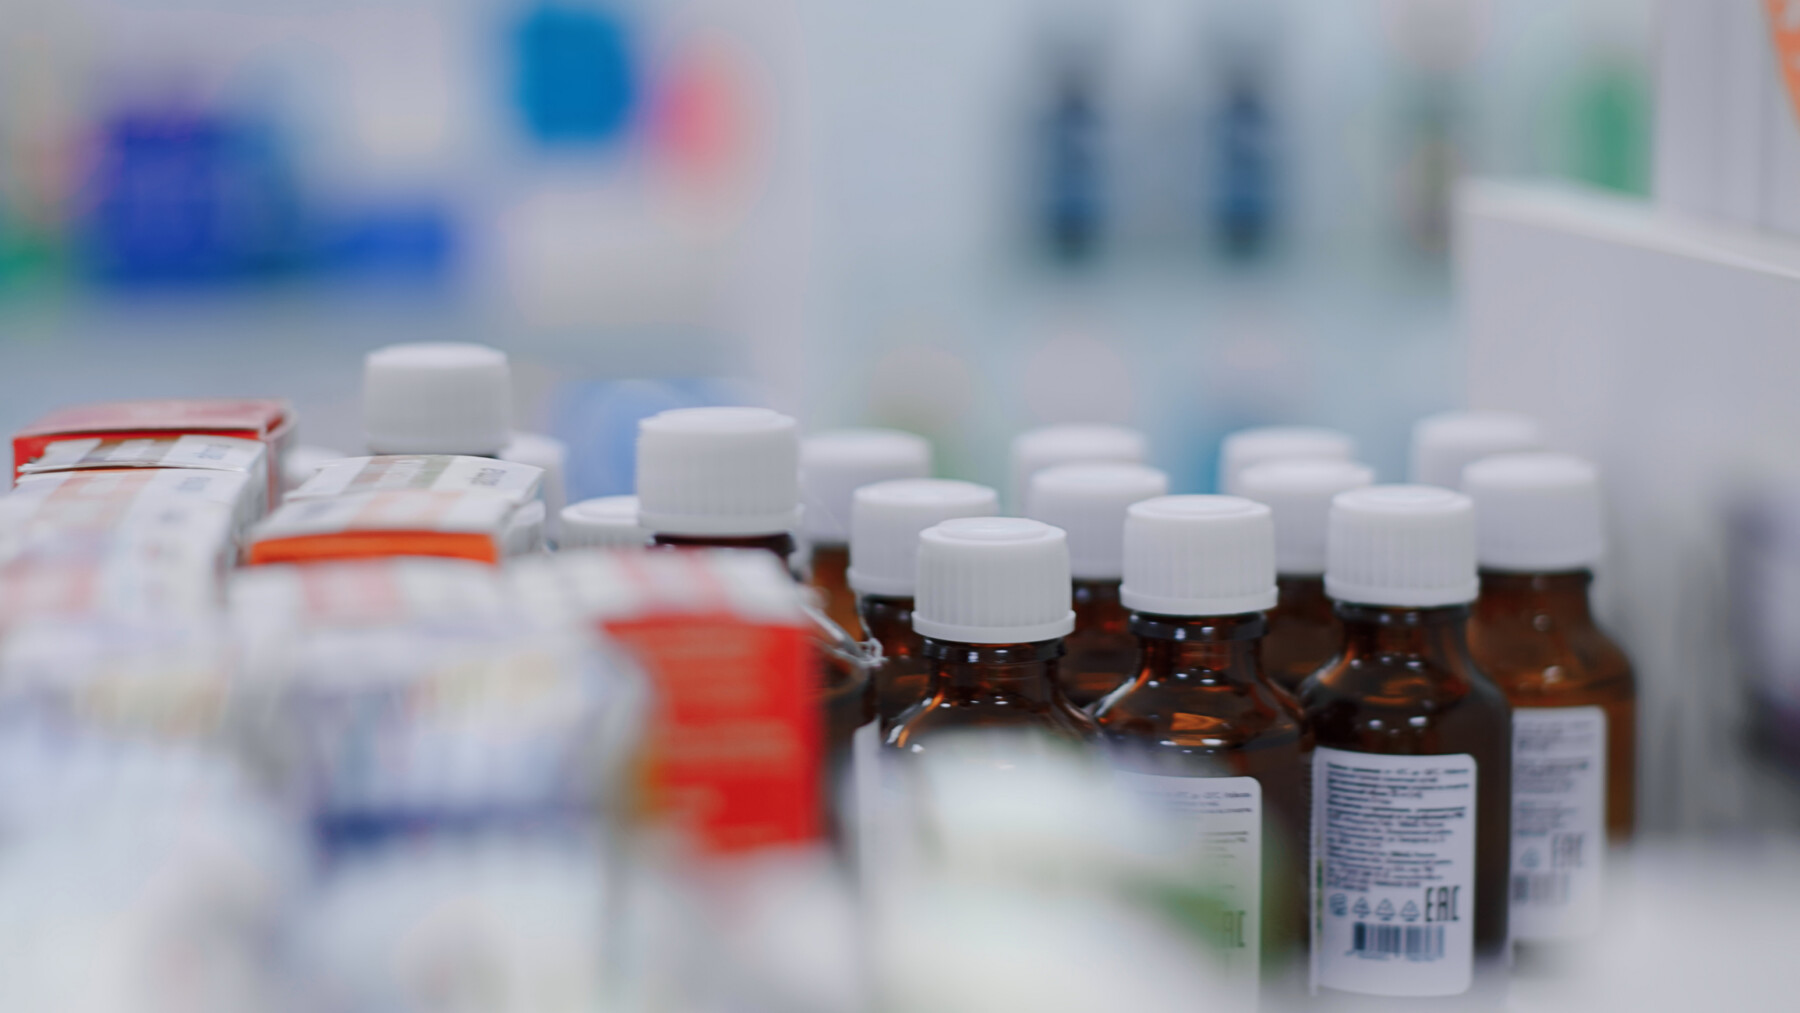
The experimental inhaled therapy ETD001 aims to ease respiratory symptoms in people with cystic fibrosis. (Image from iStock)

An experimental inhaled therapy that aims to ease respiratory symptoms in people with cystic fibrosis (CF) has won orphan drug status in the European Union — a designation designed to speed the development of new medications for rare diseases.
An arm of the European Medicines Agency (EMA) granted this status — specifically, orphan medical product designation — to ETD001, a treatment candidate being developed by Enterprise Therapeutics. It provides extra incentives to companies investing in treatments for rare diseases, defined as conditions affecting fewer than 1 of every 2,000 people in the EU. The designation gives Enterprise access to several regulatory perks, most notably a guaranteed decade of market exclusivity, free from generic competitors, if ETD001 is ultimately approved in the EU.
“Orphan drug designation represents an important regulatory milestone for the ETD001 programme and for Enterprise Therapeutics,” John Ford, PhD, the company’s CEO, said in a press release. “The decision by the EMA reflects the continued need for additional treatment options for people with cystic fibrosis.”
Ford particularly noted the need for new meds for “those who are not able to benefit from CFTR modulators,” which work to normalize mucus production. That’s the goal of ETD001, which showed positive results in a mid-stage clinical trial.
Annabella Amatulli, head of regulatory affairs at Enterprise, said the designation “further validates the strength of the programme’s scientific rationale and outcomes of the Phase 2 clinical trial,” which was completed in 2025.
“Our regulatory strategy supports our global development plan for ETD001, aligned with our goal to provide improved therapeutic options for people with cystic fibrosis,” Amatulli said.
Inhaled therapy works differently than CFTR modulators
A genetic disease, CF is caused by mutations in the gene that encodes the CFTR protein. Without a working version of this protein, the body produces unusually thick and sticky mucus, which builds up in the lungs and other organs, driving disease symptoms.
While a number of CFTR modulators — medicines that can increase CFTR activity to normalize mucus production — are approved to treat CF, these drugs only work in patients with certain underlying mutations. Further, some individuals with CF can’t take modulators due to tolerability issues.
ETD001 is an inhaled therapy designed to block the activity of ENaC (epithelial sodium channel), a different protein that also helps regulate mucus production. By inhibiting ENaC, the therapy aims to help normalize mucus production in the lungs, thus easing respiratory symptoms of CF.
Because this approach doesn’t target the mutated CFTR protein, it’s expected to work equally well in all CF patients irrespective of the underlying mutation.
A Phase 2 trial (NCT06478706) tested ETD001 in people with CF who were not taking modulators. Its results, announced earlier this year, showed the therapy was tolerated well and hinted that it may improve measures of lung function.
“We look forward to continuing the development of ETD001 in CF as well as other mucus congestion related respiratory diseases,” Ford said.

Leave a comment
Fill in the required fields to post. Your email address will not be published.